 mdi Consultants, Inc
06
Aug
FDA Releases FY 2027 MDUFA Fees
FDA Releases FY 2027 MDUFA Fees New 510(k), De Novo, PMA & Establishment Registration Costs Are Here! Is Your Regulatory Budget Ready? Effective Date: October 1, 2026 – September 30, 2027 The U.S. Food and Drug Administration (FDA) has published the Medical Device User Fee (MDUFA) rates for Fiscal Year (FY) 2027. The new fees

05
Aug
Complaint Handling Under QMSR: Where the FDA Audit Trail Begins
Short answer: Since February 2, 2026, complaint handling under 21 CFR Part 820 is governed by ISO 13485:2016 Clause 8.2.2 together with FDA's supplemental record requirements in § 820.35. Medical Device Reporting under 21 CFR Part 803 did not change: manufacturers must report a death, serious injury, or reportable malfunction within 30 calendar days of
04
Aug
FDA Warning Letter on AI-Generated cGMP Records: What It Means
FDA Issues First Warning Letter Addressing AI-Generated Drug Specifications and Master Production Records FDA issues a Warning Letter addressing “the use of AI in creating the drug product specifications, procedures, and master production or control records” The FDA recently issued a warning letter to a US pharma company that had many FDA cGMP issues. This is the first time we have seen
14
Jul
FDA Registration Renewal Opens October 1
FDA Registration Renewal Opens October 1: Your Complete Prep Checklist Every FDA-registered medical device establishment must renew its Establishment Registration between October 1 and December 31, 2026. There is no grace period. If your registration is not renewed by the deadline, it will expire on January 1, 2027, potentially resulting in shipment delays, import holds,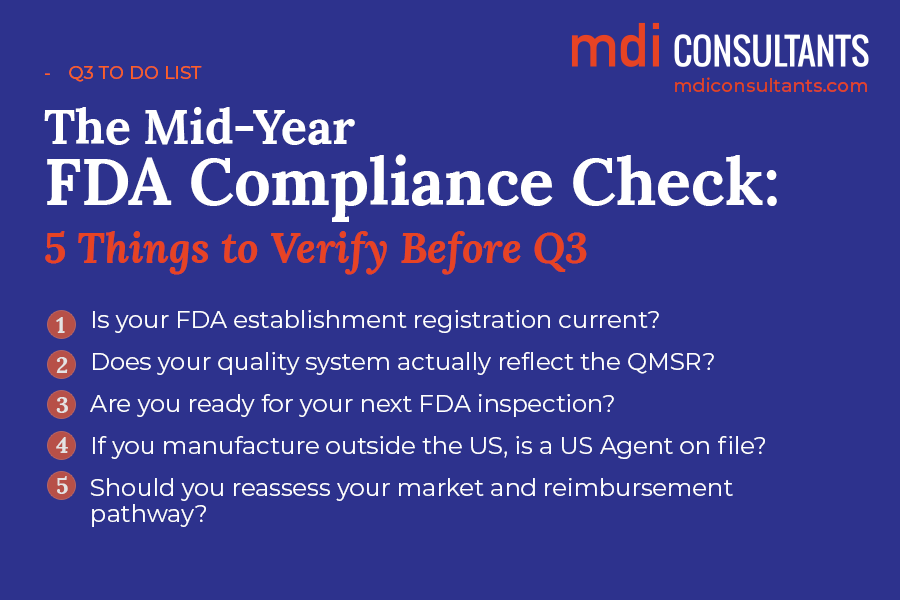
The Mid-Year FDA Compliance Check: 5 Things to Verify Before Q3
Short answer: Before Q3 begins, medical device companies should verify five key compliance areas: FDA establishment registration, QMSR implementation, inspection readiness, U.S. Agent designation (for foreign manufacturers), and the status of any open FDA 483 corrective actions. Addressing these items now can help avoid compliance issues and last-minute challenges during the October registration renewal period.

16
Jun
FDA Establishment Registration for Medical Devices — What Foreign Manufacturers Get Wrong
For foreign medical device firms, FDA establishment registration and U.S. Agent designation are not optional—they are mandatory requirements for accessing the U.S. market. Errors in either process can lead to shipment delays, import holds, or FDA Import Alerts that disrupt business operations and market access. Drawing on nearly 50 years of regulatory experience and

10
Jun
What FDA Inspectors Are Actually Checking Under the New QMSR — MDI’s First-Hand Findings
What FDA Inspectors Are Actually Checking Under the New QMSR — MDI’s First-Hand Findings The single biggest focus area in every QMSR inspection that MDI has participated in is not your quality manual, SOPs/work instruction, documentation structure or your terminology — it is your organizations ability to demonstrate risk-based thinking embedded throughout your entire quality

09
Jun
First New UV Filter Approved in Over 20 Years!
FDA Opens the Door to Sunscreen Innovation: First New UV Filter Approved in Over 20 Years! The U.S. Food and Drug Administration (FDA) has finalized an administrative order adding bemotrizinol to the Over-the-Counter (OTC) Sunscreen Monograph, making it the first new sunscreen active ingredient approved for inclusion in the U.S. OTC sunscreen framework in more

08
Jun
How to Respond to an FDA 483 and Warning Letter – Precautionary Measures
The FDA themselves state it is industry’s responsibility to work with investigators before they leave the company to remove all incorrect points on the 483. The FDA is also supposed to annotate the 483 concerning your corrective actions that were accomplished during the inspection. From latest experience with the new FDA investigators, this advice doesn’t